
TSS enrichment profile
TSS-centered enrichment provides a project-level view of whether accessibility signal is concentrated around transcription start sites before downstream peak interpretation.
ATAC Sequencing
Assays for Transposase-Accessible Chromatin using sequencing, short for ATAC-Seq, is a next-generation sequencing (NGS)-based method to study the open regions of chromatin on a genome-wide scale. ATAC-Seq has the advantages of easy operation, no crosslinking required, high signal-to-noise ratio, and low requirement for sample amount. Combined the data of ATAC-Seq, RNA-Seq, and epigenomics, the relationship between gene differential expression and regulation can be studied more accurately and comprehensively.
- Genome-wide profiling of accessible chromatin
- ATAC-seq library preparation and high-throughput sequencing
- Peak calling, annotation, differential analysis, and pathway interpretation

OverviewFeaturesWorkflowBioinformaticsDemoSamples
Overview
The DNA of eukaryotes is tightly bound to a group of proteins called histones, which constitute half the mass of a eukaryotic chromosome. There are numerous units that DNA enwinds histone formed a bead like structure called nucleosome, and the chromatin looks like many beads on a string. DNA replication and gene transcription require the unwinding of the higher structure of DNA. Open chromatin regions are these folded regions that contain important binding sites for transcription factors and transcriptional regulatory elements. ATAC-Seq utilizes the hyperactive mutant Tn5 transposase to insert sequencing adapters into accessible DNA regions, through which the open chromatin sequence can be separated from the entire chromatin, followed by next-generation sequencing and corresponding bioinformatics analysis. The research on open chromatin addresses the needs for deeper understanding of gene regulation, transcription factors that relate to cell physiology and diseases, etc.

Features
| High Throughput | Genome-Wide | Bioinformatics Analysis | One-stop Service |
|---|---|---|---|
| Cost-effective transcriptional profiling solution for large sample-size assays. | Profile all open chromatins, known and novel. | Our integrated bioinformatics pipeline can be tailored to suit your project. | Provides one-stop service for library construction, sequencing, sample QC and data analysis. |
Project Workflow
1. Sample Preparation
DNA purification;
quality assessment and quantification.
2. Library Preparation
DNA fragmentation;
library preparation.
3. Sequencing
Illumina HiSeq;
PE 50/75/100/150.
4. Data Analysis
Visualize and preprocess results, and perform custom bioinformatics analysis.
Bioinformatics Analysis Pipeline
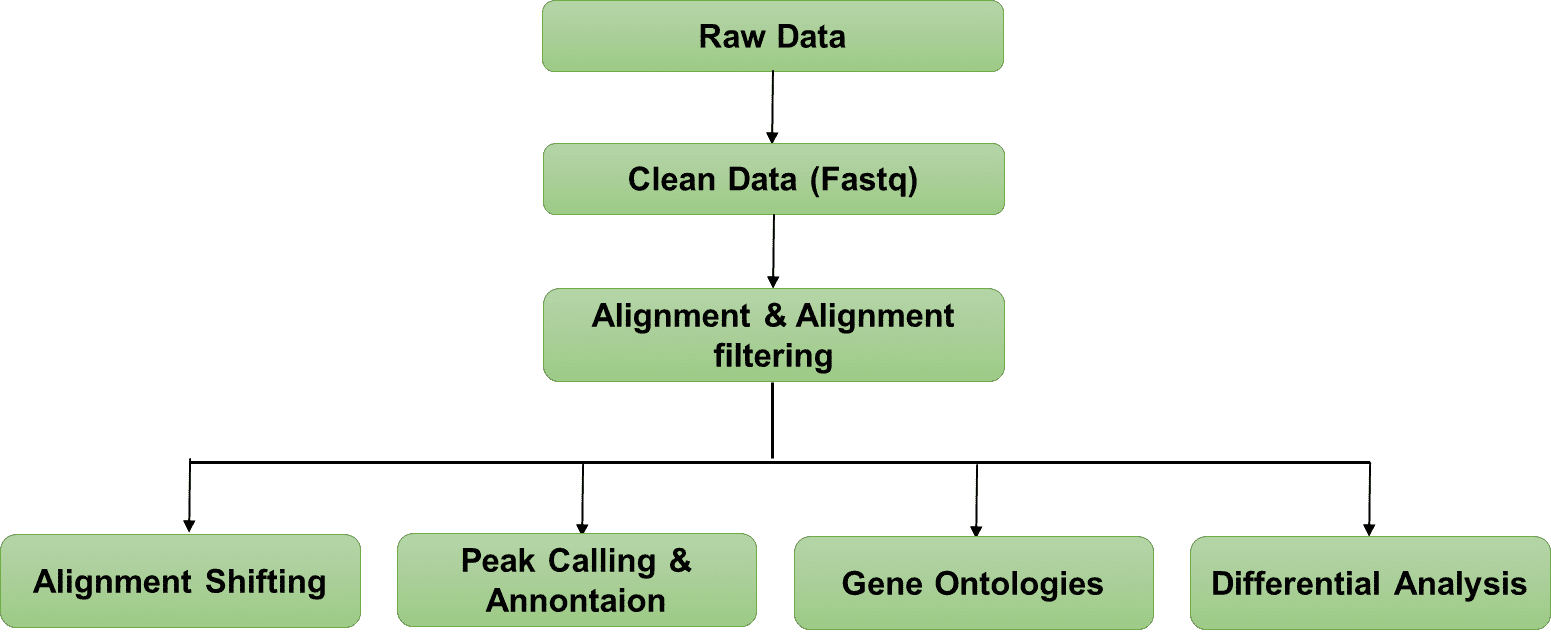
In-depth data analysis:
- Data quality control;
- Peak calling and annotation;
- Fragment size distribution;
- Gene ontologies;
- KEGG pathways;
- Differential analysis.
Representative ATAC-Seq Analysis Outputs
The following de-identified outputs from a completed bulk ATAC-seq project illustrate how data quality, peak features, between-sample patterns, differential accessibility, and functional context are evaluated together.
12 outputs · 5 stages
1
Data quality assessment
2
Peak landscape and genomic context
Peak annotation
Called peaks can be assigned to promoters, untranslated regions, exons, introns, downstream regions, and distal intergenic regions to summarize the genomic distribution of accessible chromatin.
Chromosome-wide peak distribution
Chromosome-wide peak distribution provides a genome-level overview of how accessible regions are distributed across the reference assembly.
Peak distribution relative to TSS
Peak distances relative to transcription start sites help summarize the regulatory context of accessible chromatin regions.
3
Cross-sample exploration
Peak-set overlap
Peak-set overlap analysis summarizes shared and dataset-specific accessible regions across multiple samples.
Global sample PCA
Peak-count profiles can be compared by principal component analysis to review global sample relationships, replicate consistency, group separation, and potential outliers before differential testing.
4
Differential accessibility
Differential-accessibility volcano plot
Differential peak analysis identifies regions with altered accessibility across predefined comparisons.
DDifferential-accessibility MA plot
MA plots provide a complementary view of accessibility changes across the ranked peak-signal range.
5
Functional interpretation
GO enrichment
GO enrichment analysis summarizes biological-process, cellular-component, and molecular-function patterns associated with peak-linked genes.
GO functional classification
GO functional classification organizes peak-linked genes into broad ontology categories for a complementary overview of annotation composition.
KEGG enrichment overview
KEGG dot plots compare relative enrichment, gene representation, and adjusted significance across ranked pathway categories.
KEGG pathway summary
KEGG pathway summaries provide an alternative ranked view of pathway-associated gene representation and significance.
Sample Requirements
Cell sample ≥ 50,000 cells; Nucleus sample ≥ 50,000 nuclei
Sample storage: Nuclei can be frozen at –20˚C for up to two days, avoid repeated freezing and thawing.
Shipping Method: Samples are supposed to stored in a 1.5 mL microcentrifuge tube, sealed with sealing film. Shipments are generally recommended to contain 5-10 pounds of dry ice per 24 hours.
Deliverable: FastQ, BAM, coverage summary, QC report, full statistical analysis & alignments, custom analysis reports on customer request.
References:
- Satpathy AT, Saligrama N, Buenrostro JD, et al. Transcript-indexed ATAC-seq for precision immune profiling. Nature medicine. 2018;24(5):580-90.
- Divate M, Cheung E. GUAVA: a Graphical User Interface for the Analysis and Visualization of ATAC-seq Data. Frontiers in genetics. 2018;9:250.
- Corces MR, Trevino AE, Hamilton EG, et al. An improved ATAC-seq protocol reduces background and enables interrogation of frozen tissues. Nature methods. 2017;14(10):959-62.
- Buenrostro JD, Wu B, Chang HY, Greenleaf WJ. ATAC‐seq: a method for assaying chromatin accessibility genome‐wide. Curr Protoc Mol Biol. 2015;109(1):21.9. 1-.9. 9.
* For Research Use Only. Not for use in diagnostic procedures.
×
![]()


Inquiry
Request a Demo Report
Interested in seeing a sample project report? Complete the form below and our team will send the requested demo report to your email.